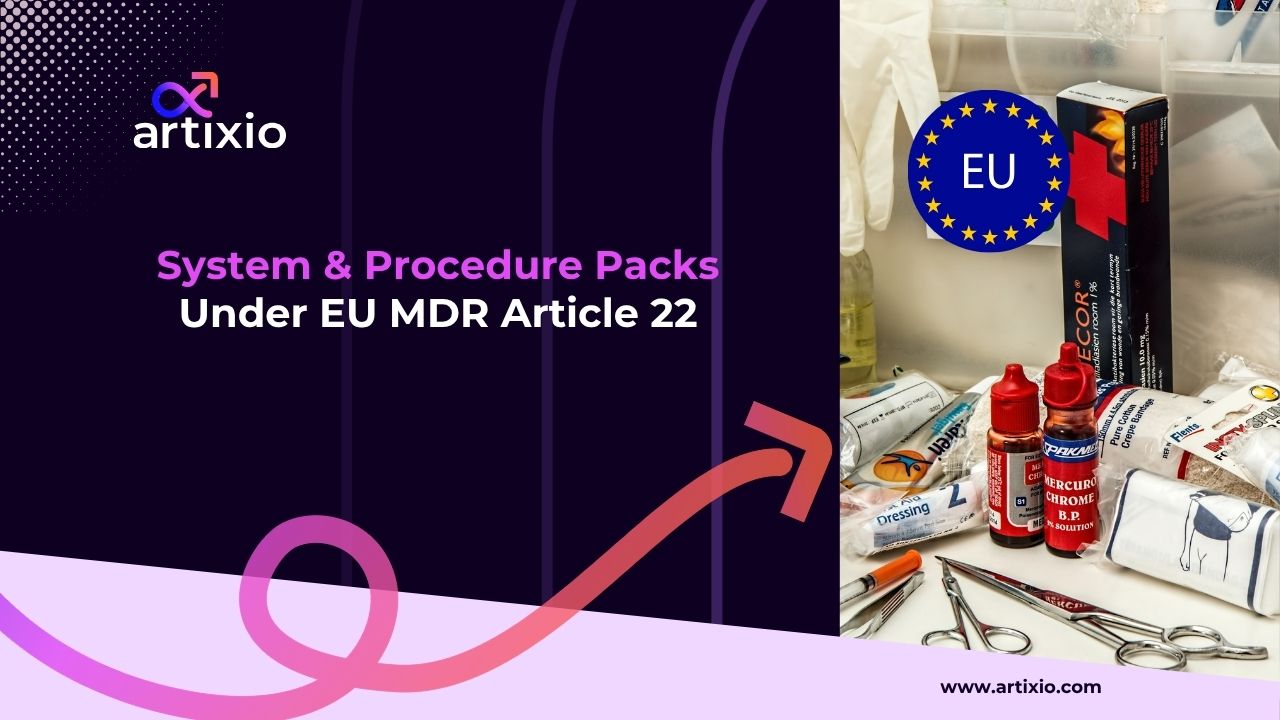
A system or procedure pack is a physical combination of medical devices and other products brought together for a specific medical purpose or procedure. A procedure pack might include things like dressings, syringes, gloves or drapes that are used together during the same treatment.
Under the EU MDR 2017/745, Article 22 covers how these systems and procedure packs can be placed on the EU market. It also explains what the company putting the pack together is responsible for.
Understanding System and Procedure Packs Under EU MDR
Under Article 2(10) of the EU MDR, a procedure pack is a combination of products packaged together and placed on the market to be used for a specific medical purpose. These packs may contain medical devices, accessories and, where justified, other products needed for the procedure. Examples can include first-aid kits, catheterization kits, wound-care packs and orthodontic procedure packs.
Article 2(11) defines a system as a combination of products, packaged together or not, that are intended to be interconnected or combined to achieve a specific medical purpose.
The main difference is simple: a procedure pack is supplied together as a pack, while a system may consist of separate products that are intended to work together for one medical purpose.
Components of System and Procedure Packs Under EU MDR
Under Article 22(1), a system or procedure pack may combine:
- CE-marked medical devices, when they are used within their intended purpose and according to the manufacturers’ instructions
- CE-marked in vitro diagnostic medical devices, where applicable and compliant with the IVDR
- Other products, including non-medical products, provided they comply with the legislation that applies to them and their inclusion is justified for the intended medical procedure
The important point is that the products cannot simply be grouped together arbitrarily. The producer must verify that the combination is compatible and that each device is being used as intended by its manufacturer.
If the pack includes a device that is not CE marked, or if a device is used outside its intended purpose, Article 22(4) may apply. In that case, the system or procedure pack is treated as a medical device in its own right and must undergo the applicable conformity assessment.
EU MDR Article 22 Requirements for System and Procedure Packs
Article 22 sets out the main requirements that apply when systems and procedure packs are placed on the EU market. These cover areas such as CE marking, conformity assessment, the producer’s responsibilities, technical documentation, labeling, sterilization and the required Article 22 statement.
CE Marking of System and Procedure Packs
If the producer combines CE-marked devices and other permitted products in accordance with their intended purposes and manufacturers’ instructions, the producer follows the Article 22 route rather than applying a new CE mark to the pack itself.
The producer must also draw up the statement required under Article 22(2), confirming that:
- the mutual compatibility of the devices and other products has been verified
- the combination has been carried out according to the manufacturers’ instructions
- the relevant information has been provided to users
- appropriate methods of internal monitoring, verification and validation have been applied
However, if the combination includes a device that does not bear the required CE marking, or a device is used outside its intended purpose, Article 22(4) applies. In that case, the system or procedure pack is treated as a device in its own right and must undergo the applicable conformity assessment under the MDR.
Conformity Assessment of System and Procedure Packs
A system or procedure pack that meets the conditions of Article 22 does not normally require a separate conformity assessment as a medical device.
However, Article 22(4) applies if:
- the pack contains devices that do not bear the required CE marking
- the devices are combined in a way that is not compatible with their intended purpose
- sterilization is carried out outside the conditions or instructions specified by the device manufacturer
In these situations, the entire system or procedure pack is treated as a medical device in its own right. The person assembling the pack then takes on the obligations of a manufacturer and must complete the applicable conformity assessment procedure under Article 52 of the MDR.
System and Procedure Pack Producer vs. Manufacturer
Under Article 22, a system or procedure pack producer is the person or company that combines CE-marked medical devices, and where applicable other compliant products, and places the resulting system or procedure pack on the EU market.
This role is different from that of a medical device manufacturer. When the producer meets the conditions of Article 22, the producer follows the specific obligations for systems and procedure packs rather than the full manufacturer obligations that apply to a device in its own right.
However, if Article 22(4) applies — for example, because the pack contains a device without the required CE marking or a device is used outside its intended purpose — the system or procedure pack is treated as a medical device in its own right. The producer then assumes the obligations of a manufacturer under the MDR.
If you need clarity on EU MDR compliance support for system and procedure packs, our regulatory consultants are available to support you at every stage. Contact at info@artixio.com.
Technical File for Systems and Procedure Packs
A system or procedure pack placed on the market under Article 22 does not normally require the same full technical documentation that applies to a medical device placed on the market in its own right.
The producer must, however, draw up the statement required under Article 22. This statement confirms that the devices and other products have been checked for compatibility, combined according to the manufacturers’ instructions, and subject to appropriate internal monitoring, verification and validation.
The statement must be kept available for the competent authorities for the retention period that applies to the devices included in the system or procedure pack. Where different retention periods apply, the longest period is used.
The Statement for Systems and Procedure Packs
Under Article 22(2), the system or procedure pack producer must prepare a statement before placing the pack on the market.
The statement should confirm that:
- the compatibility of the devices and other products has been checked
- the combination has been carried out according to the manufacturers’ instructions
- the relevant information from the manufacturers has been supplied with the pack
- the producer has applied suitable methods for internal monitoring, verification and validation
The statement must be kept available for the competent authorities for the required retention period.
Labeling Requirements for System and Procedure Packs under Article 22(5)
Systems and procedure packs placed on the market under Article 22 do not bear an additional CE marking.
They should carry:
- the name, registered trade name or registered trademark of the system or procedure pack producer
- the address where the producer can be contacted
- the information required under Annex I, Section 23 of the MDR, as applicable
- the UDI carrier in accordance with the MDR UDI requirements
The system or procedure pack must also be supplied with the relevant information from the manufacturers of the devices or other products included in the pack.
Sterilization of System and Procedure Packs
If a system or procedure pack covered by Article 22(1) is sterilized before being placed on the market, Article 22(3) applies.
The person carrying out the sterilization must use one of the conformity assessment routes allowed under Annex IX or Part A of Annex XI. The notified body’s involvement is limited to the aspects needed to ensure sterility until the sterile packaging is opened or damaged.
The person must also prepare a statement confirming that sterilization was carried out according to the manufacturer’s instructions.
If sterilization is not carried out according to those instructions, Article 22(4) applies. The system or procedure pack is then treated as a medical device in its own right, and the person responsible takes on the obligations of a manufacturer and must follow the applicable conformity assessment procedure under Article 52.
Classification, UDI and EUDAMED Requirements
Systems and procedure packs also have specific classification, UDI and EUDAMED requirements. These requirements help identify the pack, record the relevant risk-class information and ensure the required data is registered correctly.
Risk Classification of Systems and Procedure Packs
Systems and procedure packs placed on the market under Article 22 are not given a separate MDR classification in the same way as individual medical devices.
However, a risk-class value is still required for UDI/EUDAMED registration. For this purpose, the highest risk class of the medical device components included in the system or procedure pack should be used.
If a pack contains Class I and Class IIa devices, Class IIa is entered in EUDAMED.
This classification value is mainly used for registration and UDI-related requirements; it does not mean that the procedure pack itself has undergone classification as a separate medical device.
EU Medical Device Classification
Under the EU MDR, medical devices are classified into four main risk classes: Class I, Class IIa, Class IIb and Class III. The classification depends on the device’s intended purpose and the rules set out in Annex VIII of the MDR.
| Class | General Risk Level |
| Class I | Lower risk |
| Class IIa | Low to medium risk |
| Class IIb | Medium to high risk |
| Class III | Highest risk |
Class I also has subcategories such as Is, Im and Ir for sterile devices, measuring devices and reusable surgical instruments.
Other factors can also affect classification, including how invasive the device is, how long it is used, whether it is active or implantable, and where it is used in the body.
Basic UDI-DI Assignment and UDI Database Registration:
System and procedure pack producers must register in EUDAMED and obtain a Single Registration Number (SRN).
Before placing a system or procedure pack on the market under Article 22(1) or 22(3), the producer must assign a Basic UDI-DI. Systems or procedure packs with the same group of components and the same intended purpose may share the same Basic UDI-DI.
Each system or procedure pack needs its own UDI. The producer then records the required UDI details in the database.
Since 28 May 2026, the relevant EUDAMED modules have been mandatory. This means system and procedure pack producers now have to enter their registration and product details in EUDAMED.
Difference Between UDI-DI and UDI-PI
- The UDI-DI identifies a specific system or procedure pack. It is linked to information such as the product reference, name and other registration details.
- The UDI-PI carries production information. Depending on the product, this may include details such as the lot or batch number, serial number, manufacturing date or expiry date.
In simple terms, the UDI-DI tells which product it is, while the UDI-PI tells which production unit or batch it belongs to.
Additional Regulatory Considerations for System and Procedure Packs
Some system and procedure pack situations need additional regulatory consideration, especially when legacy devices, non-EU producers or post-market vigilance are involved. The applicable requirements can vary depending on how the pack is assembled and placed on the market.
Utilizing MDD/AIMDD Legacy Devices in Systems and Procedure Packs
Before the EU MDR, medical devices in Europe followed older rules known as the MDD and AIMDD. Some devices certified under those older rules are still allowed on the market during the MDR transition period. These are commonly referred to as legacy devices.
Legacy devices that continue to benefit from the MDR transitional provisions may still be included in systems and procedure packs, but the applicable route depends on the combination.
If a system or procedure pack consists only of legacy devices and the required declaration under Article 12(2) of the MDD was drawn up before 26 May 2021, the transitional provisions may continue to apply. In such cases, Article 22 of the MDR does not apply during the relevant transition period.
The transition period for that legacy system or procedure pack ends when the transition period ends for the legacy device with the highest MDR risk class included in the pack.
Where legacy devices are used in a system or procedure pack that does not qualify for the specific legacy SPP transitional route, the applicable MDR requirements should be assessed for that combination. The producer should also confirm that every legacy device included can still lawfully be placed on the market under the MDR transitional provisions.
Appointing a European Authorized Representative for System and Procedure Packs
A system or procedure pack producer established outside the EU does not need to appoint an EU Authorized Representative solely because it places an Article 22 system or procedure pack on the market.
This is different from a non-EU medical device manufacturer, which must appoint an Authorized Representative in the Union.
However, if Article 22(4) applies and the system or procedure pack becomes a medical device in its own right, the producer takes on the obligations of a manufacturer. In that situation, a producer established outside the EU would also need to meet the MDR requirements that apply to non-EU manufacturers, including appointing an Authorized Representative.
The individual CE-marked devices included in the pack must, of course, already meet their own MDR requirements. If one of those device manufacturers is based outside the EU, that manufacturer is responsible for having its own Authorized Representative where required.
Handling Post-market Vigilance of System and Procedure Packs
The MDR does not set out a separate vigilance-reporting process specifically for system and procedure pack producers acting under Article 22.
If an incident appears to be linked to one of the medical devices included in the pack, the SPP producer should pass the information to the relevant device manufacturer so that it can be assessed under the manufacturer’s vigilance process. Manufacturers are responsible for reporting serious incidents and field safety corrective actions under Article 87 of the MDR.
It is therefore useful for SPP producers and device manufacturers to define how incident information will be exchanged between them.
Where an issue appears to arise from the SPP producer’s own activities, such as combining or sterilizing the products, the circumstances should be assessed carefully. If the conditions of Article 22(4) are found to apply, the system or procedure pack is treated as a device in its own right and the producer assumes the applicable manufacturer obligations.
Steps to Place Systems and Procedure Packs on the EU Market
Before placing a system or procedure pack on the EU market, the producer should:
1. Check the products being combined
Make sure the devices bear the required CE marking and are being used within their intended purpose and the limits set by their manufacturers.
2. Verify compatibility
Confirm that the devices and any other products can be combined safely and in line with the manufacturers’ instructions.
3. Prepare the Article 22 statement
The producer must document the compatibility checks, packaging and user information, and the internal monitoring, verification and validation applied to the combination.
4. Complete sterilization requirements, where applicable
If the system or procedure pack is sterilized before being placed on the market, the requirements of Article 22(3) apply, including limited Notified Body involvement for the sterility aspects.
5. Apply the required labeling and UDI
The pack should carry the producer’s name and address, the applicable information required under Annex I, and its own UDI.
6. Register the producer and the system or procedure pack in EUDAMED
The producer must obtain an SRN and complete the relevant UDI/EUDAMED registration requirements.
7. Keep the required statement available
The Article 22 statement must be retained for the applicable period and made available to competent authorities when required.
One important exception should follow the list:
If the conditions of Article 22(4) are triggered — for example, the pack includes a device without the required CE marking, the combination is incompatible with the original intended purpose, or sterilization is not carried out according to the manufacturer’s instructions — the pack is treated as a device in its own right. The person responsible then assumes the obligations of a manufacturer and must follow the applicable conformity assessment procedure under Article 52.
Conclusion
System and procedure packs can make it easier to bring together the products needed for a specific medical purpose, but Article 22 places clear responsibilities on the producer.
The main points are to check device compatibility, follow the manufacturers’ instructions, prepare the required statement, meet sterilization and labeling requirements where applicable, and complete the relevant UDI and EUDAMED registration steps.
If the conditions of Article 22 are not met, the pack may be treated as a medical device in its own right, which brings additional manufacturer and conformity assessment obligations.
Artixio’s regulatory professionals have 15–35 years of experience across global and regional regulatory requirements and can support companies with EU MDR compliance, market access and ongoing product maintenance. For assistance with system and procedure pack requirements, contact us at info@artixio.com.
FAQs
1. What is the difference between a system and a procedure pack under the EU MDR?
A procedure pack is a combination of products packaged together and placed on the market for a specific medical purpose. A system is also a combination of products intended to work together for a specific medical purpose, but the products do not necessarily have to be packaged together.
2. Do systems and procedure packs require a separate CE mark?
Not when they meet the conditions of Article 22. In that case, the system or procedure pack itself does not receive an additional CE marking. If Article 22(4) applies, the pack is treated as a medical device in its own right and the applicable manufacturer and conformity assessment requirements apply.
3. When does a system or procedure pack become a medical device in its own right?
This can happen when the conditions of Article 22 are not met, such as when a required CE marking is missing, a device is used outside its intended purpose, or sterilization is not carried out according to the manufacturer’s instructions. In that situation, the person responsible takes on the obligations of a manufacturer.
4. Does a system or procedure pack need its own UDI?
Yes. A system or procedure pack must have its own UDI, including a UDI-DI and UDI-PI. A Basic UDI-DI is also assigned for registration purposes.
5. Is a Notified Body always required for systems and procedure packs?
No. Notified Body involvement is not automatically required for every Article 22 system or procedure pack. It becomes relevant in specific situations, such as sterilization under Article 22(3), where its role is limited to the sterility aspects, or when the pack is treated as a device in its own right.
6. Can legacy MDD or AIMDD devices still be included in a system or procedure pack?
They may be included if they still qualify under the MDR transitional provisions and the relevant conditions are met. The applicable route depends on whether the pack contains only legacy devices or a combination of legacy and MDR-compliant devices.
7. Does a non-EU system or procedure pack producer need an EU Authorized Representative?
Not solely because it acts as a system or procedure pack producer under Article 22. If Article 22(4) applies and the pack becomes a medical device in its own right, the producer takes on manufacturer obligations. A non-EU manufacturer would then need to meet the applicable EU Authorized Representative requirements.