The healthcare landscape is in a perpetual state of transformation. As a consequence, there is a noticeable surge in the development of innovative combination products. They are engineered to offer groundbreaking therapies and solutions. These combination products present distinctive opportunities and challenges. That’s particular in the context of regulatory scrutiny, approval processes, and subsequent post-approval compliance requirements.
What Are Combination Products?
A combination product contains two or more regulated components, which can be drugs, devices, or biologics. There are four categories as given under 21 CFR 3.2(e) mentioned below:
- Drug and Device
- Biological Product and Device
- Drug and Biological Product
- Drug, Device, and Biological Product
These combination products may be introduced to the market. Their components might either be bundled into a single entity or individually packaged but labeled for co-use. FDA also considers some separately packaged products as combination products. That happens when their labelling or investigational plan specifies that they must be used together for the intended purpose. This can include a product intended for use only with a specific, individually identified approved or investigational product.
Regulatory Bodies
FDA’s three main medical product centers are the following:
- Center for Drug Evaluation and Research (CDER)
- Center for Biologics Evaluation and Research (CBER)
- Center for Devices and Radiological Health (CDRH)
They look after the review and regulation of drugs, biological products, and medical devices, respectively. However, some products challenge these distinctions. There may be ambiguity and blurring of the lines between them.
The FDA established the Office of Combination Products (OCP). It coordinates the regulation of combination products. OCP assigns products to the appropriate center. It coordinates reviews that involve multiple centers and oversees combination product post-market activities.
In instances where ambiguity arises, the FDA has put in place processes to resolve such issues. Also, it determines which FDA center should regulate the product. Combination product manufacturers can get clarity through the FDA’s RFD or Pre-RFD submissions. Choosing the correct regulatory path and communicating with the OCP becomes possible with these processes.
Manufacturers have the option to approach the OCP to classify their combination product and identify the lead center responsible for pre-market review submissions. The OCP utilizes two distinct processes for this purpose:
- The Pre-Request for Designation (Pre-RFD)
- The Request for Designation (RFD)
Under Section 503(g) of the Federal Food, Drug, and Cosmetic Act, FDA assigns the lead center for a combination product based on its primary mode of action (PMOA). The PMOA is the single mode of action that provides the most important therapeutic action of the combination product. If the FDA cannot determine the PMOA, it uses the statutory and regulatory assignment process. This considers the center with the most expertise to review the product’s most significant safety and effectiveness questions.
Are RFD and Pre-RFD Submissions Mandatory?
Not every combination product requires an RFD or Pre-RFD. Sponsors use these processes when the product’s classification or FDA center assignment is unclear or disputed. If the regulatory classification and responsible center are already clear, sponsors can work directly with the appropriate FDA center.
A Pre-RFD is an optional route for obtaining preliminary, non-binding feedback. An RFD provides a formal, binding determination.
Regulatory requirements and guidance can change over time. Artixio’s QuriousRI regulatory intelligence platform helps regulatory teams monitor health authority updates. Get access to expert-reviewed summaries and assess their potential impact on products and compliance activities.
Key Benefits of the RFD and Pre-RFD Processes
Here are the crucial benefits to know of the RFD and Pre-RFD processes:
Regulatory Clarity: These processes clarify which agency center will regulate the combination product.
Early Engagement: Sponsors can engage with the FDA early in the development process. They can get insights and feedback in return.
Enhanced Communication: Open communication with the FDA lets sponsors manage concerns and resolve potential issues in the beginning.
Regulatory Direction: Identifying the correct FDA classification and lead center helps sponsors plan the proper regulatory pathway.
Pre-Request for Designation (Pre-RFD)
The pre-RFD process allows sponsors to get the FDA’s preliminary, non-binding assessment of a human medical product’s classification and/or the FDA center responsible for its review. Sponsors can use this process when they are not sure about the product’s regulatory identity or center assignment. A Pre-RFD may be submitted at any stage of product development. That is, provided the sponsor has enough information for the FDA to assess the product.
There is no compulsory page limit for pre-RFD submissions. However, the FDA recommends keeping them concise. The November 2025 guidance states that the information can be presented in 15–20 pages. It excludes courtesy copies of published literature. FDA’s preliminary assessment doesn’t bind the agency and the sponsor.
Authorities review a Pre-RFD within 10 business days of receipt. They check whether it contains enough information for an assessment. After acknowledging the acceptance of the submission for review, it aims to provide feedback within 60 calendar days. The actual review may take longer if the submission doesn’t have information or raises complex issues.
A Pre-RFD response does not identify the premarket application type that a sponsor should submit. It addresses product classification and/or center assignment. Sponsors should receive consultation from the proper FDA center for advice on the premarket route.
Request for Designation (RFD)
The RFD process is established under 21 CFR Part 3. Sponsors receive a formal FDA determination of a product’s regulatory classification and/or the FDA center responsible for its review under this. They should submit an RFD before filing an investigational or marketing application. That is if the product’s classification or center assignment is unclear or disputed. RFD submissions are limited to a maximum of 15 pages. This limit excludes attachments.
RFD submissions can be initiated at any phase during the development of a medical product. Within five business days of receiving an RFD, OCP reviews it for completeness. If the submission meets the requirements, FDA confirms its filing date. FDA must issue its designation letter within 60 calendar days of that filing date. If it does not issue the letter within this period, the sponsor’s recommended classification or center assignment becomes the designation under 21 CFR 3.8(b).
21 CFR 3.7 says that an RFD must include the following:
- Sponsor’s identity
- Their contact details
- Product classification and names
- Product description
- Component information
- Composition
- Development and testing information
- Manufacturing process
- Proposed use or indications
- Modes of action
- Schedule and duration of use
- Dose and route of administration where applicable
- Information on related products
- Any other relevant details
The sponsor must also recommend which FDA center should have primary jurisdiction. FDA’s designation letter is binding on the agency and the sponsor for the product as described in the submission. However, the determination may be changed under the conditions set out in Section 563 of the FD&C Act and 21 CFR 3.9.
Information to be included in RFD and Pre-RFD Submissions
The fundamental elements to be incorporated in RFD and Pre-RFD applications consist of
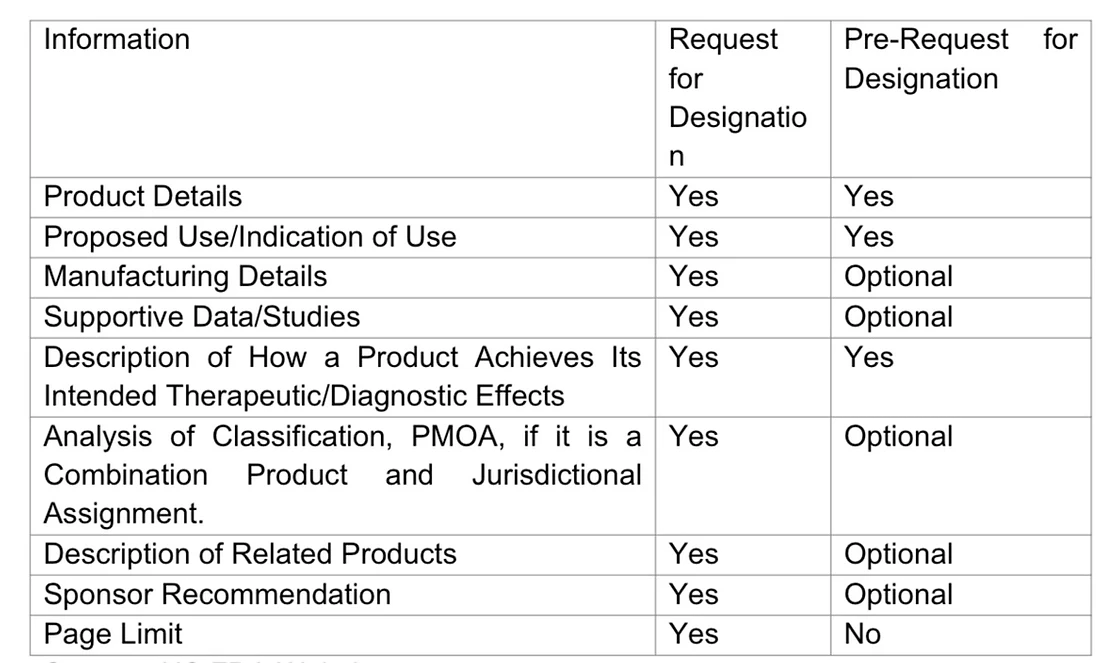
Source: US FDA Website
Information to Include in a Pre-RFD Submission
FDA’s November 2025 guidance recommends that sponsors provide enough information for OCP to understand the product. Also, so it can assess its classification and center assignment. The submission should include the following:
- Sponsor’s name, address, and contact details.
- Product name and a complete description of the product.
- Names, sources, and regulatory history of all constituent components.
- Product photographs or diagrams.
- Proposed intended use, indications, and claims.
- Instructions for use and conditions of use.
- Full active and inactive components list. It includes their composition and quantities or concentrations.
- Details of biologically derived materials. It covers their sources and processing methods, where applicable.
- An explanation of how the product and its components achieve the intended purpose.
- Available information on the relative contribution of each component to the product’s therapeutic or diagnostic effects, including the PMOA where relevant.
- An explanation of how the product will be marketed, including whether its components will be combined, co-packaged, or sold separately for use together.
Information to Include in an RFD Submission
Under 21 CFR 3.7, an RFD must contain the information FDA needs to make a formal classification and jurisdictional determination. The submission should cover the following:
- Sponsor’s name, address, establishment registration number (where applicable), and contact details
- Product classification and the common, generic, or usual name of the product and its components
- Product’s proprietary name, if available
- Details of any component that has received premarket approval, an investigational exemption, or another FDA regulatory determination
- Chemical, physical, or biological composition
- Development history and testing results, including animal testing
- Manufacturing processes and sources of the components
- Proposed use or indications
- All known modes of action and the sponsor’s PMOA analysis
- Schedule and duration of use
- Dose and route of administration for drug or biological components
- Details of related products and their regulatory status
- Other information relevant to the FDA’s determination
- Sponsor’s recommendation on which FDA center should have primary jurisdiction
Conclusion
Combination products hold great promise for improving patient care and advancing healthcare. However, their regulatory complexity can be a challenge for product developers. The Request for Designation (RFD) and Pre-RFD processes by the US FDA are essential tools. They can ensure proper evaluation and regulation of these innovative products. Early engagement with the FDA can help sponsors follow rules more effectively.
Ready to determine the right FDA regulatory pathway for your combination product? Artixio provides US FDA regulatory and compliance services to help manufacturers navigate regulatory requirements and bring their products to the US market. Contact our experts at info@artixio.com for guidance on your combination product regulatory needs.
FAQs
1. Can a sponsor submit a Pre-RFD if the product’s classification is already clear?
Yes. A Pre-RFD is optional. It can provide preliminary, non-binding feedback on product classification or FDA center assignment. A sponsor can work directly with that FDA center if the classification and responsible center are clear.
2. Does an FDA Pre-RFD response determine which premarket application a sponsor must file?
No. A Pre-RFD response addresses classification and/or center assignment. It does not identify the required premarket application; sponsors should consult the appropriate FDA center about the premarket pathway.
3. What happens if the FDA does not issue an RFD designation letter within 60 calendar days?
If the FDA does not issue the designation letter within 60 calendar days of the RFD filing date, the sponsor’s recommended classification or center assignment becomes the designation. It is described in 21 CFR 3.8(b). The determination applies to the product as described in the submission and may be changed under the conditions in Section 563 of the FD&C Act and 21 CFR 3.9.