The Electronic Common Technical Document (eCTD) is the standard electronic format used for many regulatory submissions involving drugs and biologics.
It organizes CTD content in a standard structure so regulatory authorities can review and manage submissions electronically.
This article explains the eCTD format, the five-module CTD structure, and the submission processes followed by three major regulatory authorities: the FDA, EMA and TGA.
What Is eCTD?
An eCTD submission is built as a structured electronic dossier rather than as a collection of unrelated files. The format helps regulators locate information quickly and also makes it easier to manage updates submitted later in the product lifecycle.
One of the key features of eCTD is lifecycle management. New documents, replacements and updates can be linked to earlier submission content, so reviewers can see how the dossier changes over time.
eCTD Regulatory Requirements
Each authority has its own technical rules for eCTD submissions. These cover areas such as the accepted eCTD version, regional Module 1 requirements, file formats, controlled terminology and validation criteria.
The details vary by region. FDA currently supports both eCTD v3.2.2 and v4.0. Since September 16, 2024, eCTD v4.0 has been accepted for new NDA, BLA, ANDA, IND and master file submissions to CDER and CBER.
Existing applications created in v3.2.2 continue in that format until FDA enables forward compatibility.
Structure And Components Of eCTD
The CTD is organized into five modules. Module 1 is region-specific, while Modules 2, 3, 4 and 5 are intended to be common across regions. Each module is discussed below.
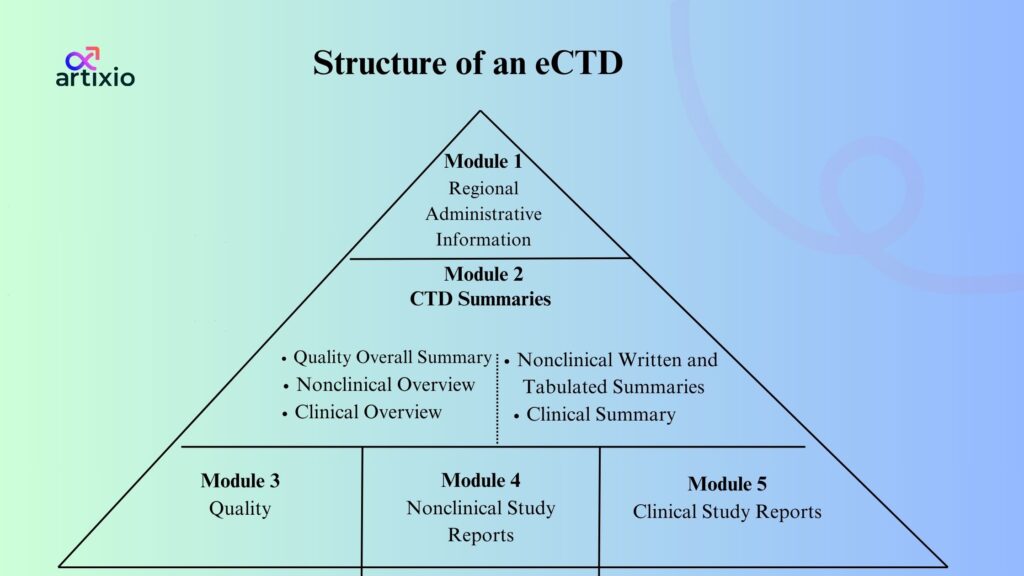
Module 1: Regional Administrative Information
Module 1 contains information that is specific to the region where the submission is being made. This can include application forms, prescribing information and other administrative documents required by the relevant regulatory authority.
Module 1 is not part of the harmonized CTD. Its content and format are defined by each region.
Module 2: Common Technical Document Summaries
Module 2 brings together the key summaries and overviews for the quality, nonclinical and clinical information presented in the later modules.
It includes:
- 2.1 CTD Table of Contents
- 2.2 CTD Introduction
- 2.3 Quality Overall Summary
- 2.4 Nonclinical Overview
- 2.5 Clinical Overview
- 2.6 Nonclinical Written and Tabulated Summaries
- 2.7 Clinical Summary
Module 3: Quality
Module 3 contains the quality information for the drug substance and drug product, including chemistry, manufacturing and controls.
The main sections are:
- 3.1 Table of Contents of Module 3
- 3.2 Body of Data
3.2.S Drug Substance
3.2.P Drug Product
3.2.A Appendices
3.2.R Regional Information - 3.3 Literature References
Module 4: Nonclinical Study Reports
Module 4 contains the nonclinical study reports used to support the safety assessment of a pharmaceutical product.
The main sections are:
- 4.1 Table of Contents of Module 4
- 4.2 Study Reports
Pharmacology
Pharmacokinetics
Toxicology - 4.3 Literature References
Module 5: Clinical Study Reports
Module 5 contains the clinical study reports and related information used to support the efficacy and safety evaluation of a pharmaceutical product.
The main sections are:
- 5.1 Table of Contents of Module 5
- 5.2 Tabular Listing of All Clinical Studies
- 5.3 Clinical Study Reports
- 5.4 Literature References
FDA Submission Types That Require eCTD
For submissions to FDA’s CDER and CBER, the eCTD requirement applies to:
- New Drug Applications (NDAs)
- Abbreviated New Drug Applications (ANDAs)
- Biologics License Applications (BLAs)
- Commercial Investigational New Drug applications (INDs)
- Subsequent submissions to these application types, including amendments, supplements and reports
- Master files, such as Drug Master Files (DMFs), when they are subject to the eCTD requirement
FDA also accepts eCTD for some other submission types, but the requirement is not the same in every case. For example, eCTD is optional but encouraged for non-commercial INDs, submissions for blood and blood components, and Type III master files.
FDA eCTD Submission Process
For submissions to FDA, the process generally involves the following steps:
1. Obtain the applicable application or submission number
Where required, obtain the appropriate pre-assigned application or submission number before transmitting the eCTD sequence.
2. Prepare the eCTD according to the supported FDA specifications
The submission should follow the applicable eCTD version, FDA regional Module 1 requirements and current validation criteria.
3. Set up access to ESG NextGen
FDA uses the Electronic Submissions Gateway Next Generation (ESG NextGen) as the secure entry point for electronic regulatory submissions.
4. Submit the eCTD package through ESG NextGen
Submissions can be sent through the Unified Submission Portal, API integration or other supported ESG NextGen connection methods.
5. Review FDA acknowledgements and validation results
After transmission, FDA checks the submission for basic technical requirements and then applies eCTD validation rules. Any validation errors should be reviewed and corrected where required.
EMA eCTD Submission Process
For human medicines submitted through the centralised procedure, EMA requires eCTD submissions to be sent through the eSubmission Gateway or eSubmission Web Client.
The process generally includes:
1. Prepare the dossier in eCTD format
The submission should follow the applicable EU eCTD specifications and the current regional Module 1 requirements.
2. Prepare the required delivery information
The correct submission metadata and XML delivery file should be included based on the procedure and submission type.
3. Submit through the eSubmission Gateway or Web Client
These are the mandatory electronic submission channels for centralised human medicinal product applications.
4. Check the acknowledgement and technical validation feedback
After submission, the system provides confirmation of receipt and technical validation feedback. Any technical issues should be reviewed and corrected where necessary.
TGA eCTD Submission Process
TGA requires prescription medicine applications, including master files, to be submitted in eCTD format. Other specified product categories may also use eCTD.
For Australia, eCTD submissions must follow the current TGA regional requirements for Module 1 together with the applicable ICH specifications for Modules 2–5.
The process generally includes:
- Confirm that eCTD is the accepted format for the regulatory activity.
- Prepare the Australian Module 1 content using the current TGA specification.
- Compile the remaining dossier content according to the applicable ICH eCTD structure.
- Publish and validate the eCTD sequence before submission.
- Submit the sequence through the TGA’s accepted electronic submission process.
Advantages Of eCTD
eCTD reduces the need for large paper-based regulatory dossiers and makes submissions easier to store, access and review electronically.
It also improves navigation within the dossier. Reviewers can locate information more easily, including documents submitted with the original application and information added later in the product lifecycle.
Other advantages include:
- a standardized submission structure;
- easier access to regulatory information;
- support for lifecycle management and future updates;
- less duplication when information can be reused or referenced;
- easier review across large regulatory dossiers.
Challenges in eCTD Submissions
Preparing an eCTD submission can involve technical and operational challenges, especially when companies are working across different regulatory regions. Some common challenges include:
Meeting technical validation requirements: Errors in the submission structure, XML, file formats or metadata can affect whether a sequence passes technical validation.
Regional differences: Module 1 requirements, controlled terminology, validation criteria and accepted technical specifications can vary between regulatory authorities.
Managing the submission lifecycle: Each new sequence must be prepared in a way that correctly relates to the documents and regulatory activity submitted earlier.
Keeping up with specification updates: Authorities periodically update validation rules, regional specifications and supported eCTD versions, so submission teams need to work with the current requirements.
Transitioning between eCTD versions: The shift from eCTD v3.2.2 to v4.0 is happening at different stages in different regions, which can make global submissions more complicated.
How to Manage eCTD Submission Challenges
Companies can reduce eCTD submission issues by focusing on a few practical areas:
Use the latest technical specifications: Check the current regional Module 1 requirements, validation criteria, controlled terminology and supported eCTD version before publishing a sequence.
Validate before submission: Run the sequence through appropriate validation tools and correct technical errors before sending it to the regulatory authority.
Maintain lifecycle consistency: Make sure new sequences correctly relate to earlier submission content and use the appropriate lifecycle operations.
Manage regional requirements separately: FDA, EMA and TGA each have their own regional specifications and validation rules, even though the core CTD structure is harmonized.
Keep publishing tools and internal procedures updated: Changes to eCTD specifications, validation criteria and controlled vocabularies should be reflected in the software and processes used to prepare submissions.
Conclusion
eCTD provides a structured way to prepare, submit and manage regulatory information throughout the product lifecycle. The five-module CTD structure helps organize quality, nonclinical, clinical and regional administrative information, while eCTD lifecycle management makes later updates easier to track.
However, the technical requirements are not identical across regions. Companies need to follow the current eCTD version, regional Module 1 specifications, validation criteria and submission process required by the relevant authority.
Need support with preparing or submitting an eCTD dossier? Explore Artixio’s eCTD publishing and submission services for support with submission planning, publishing, lifecycle submissions, dispatch and troubleshooting.
FAQs
1. What is the full form of eCTD?
eCTD stands for Electronic Common Technical Document. It is the electronic format used to organize and submit CTD content to regulatory authorities.
2. What are the five eCTD modules?
The eCTD follows the CTD module structure:
- Module 1: Regional administrative information
- Module 2: CTD summaries
- Module 3: Quality
- Module 4: Nonclinical study reports
- Module 5: Clinical study reports
Module 1 is region-specific, while Modules 2–5 follow the harmonized CTD structure.
3. Why is eCTD validation important?
Validation checks whether the submission meets the technical requirements of the relevant authority. Errors in areas such as XML, file formats, metadata or dossier structure can prevent a submission from passing technical validation and may need to be corrected before processing.
FDA, for example, maintains separate current validation criteria for eCTD v3.2.2 and v4.0 and updated both sets in August 2026.
4. Is eCTD v4.0 mandatory for all FDA submissions?
No. FDA currently supports both eCTD v3.2.2 and v4.0. FDA currently accepts new NDA, BLA, ANDA, IND and master file applications in eCTD v4.0, while existing v3.2.2 applications continue in that format until FDA implements forward compatibility.
5. Is Module 1 the same in every country?
No. Module 1 is region-specific. Each regulatory authority sets its own administrative and technical requirements for this module. Modules 2–5 follow the common CTD structure.
6. Can a company manage eCTD submissions in-house?
Yes. Companies can do eCTD submissions on their own. They just need staff who understand the process and the right software. External help may still be useful when the work becomes too much or when a submission is for a new market.