
PADER is short for Periodic Adverse Drug Experience Report. It is one of the ways FDA receives periodic postmarketing safety information for approved drugs. Under 21 CFR 314.80(c)(2), certain applicants must submit these reports at defined intervals.
The report brings together adverse drug experience information from that period and includes the required summary and analysis. It is used after approval to help FDA keep track of postmarketing safety information and any actions taken because of adverse drug experiences.
In this article, we explain how PADER works, what information goes into the report, when it has to be submitted, and how it differs from PSUR and PBRER.
What Is a Periodic Adverse Drug Experience Report (PADER)?
A PADER is a periodic safety report submitted to the U.S. FDA for approved drug applications. Under 21 CFR 314.80(c)(2), it covers adverse drug experiences that were not already reported through the 15-day Alert process.
A PADER also gives FDA an overview of safety information collected during the reporting period. It includes a narrative summary and analysis of the adverse drug experience information, an analysis of 15-day Alert reports already submitted during that interval, and a history of actions taken because of adverse drug experiences, such as labelling changes or additional studies.
Serious and unexpected adverse drug experiences that meet the expedited-reporting criteria are handled separately through 15-day Alert reports rather than waiting for the next PADER.
Why Is PADER Important For Drug Safety?
PADER is important because it gives FDA a periodic view of adverse drug experience information collected after a drug is marketed. The report brings together safety information from the reporting period and helps FDA review whether any patterns or changes need further attention.
The report also includes a summary and analysis of the information submitted during the interval, an analysis of 15-day Alert reports, and a history of actions taken because of adverse drug experiences, such as labelling changes or additional studies.
Regular periodic reporting also helps applicants meet their postmarketing safety-reporting obligations under 21 CFR 314.80(c)(2) and maintain an ongoing record of safety information after approval.
PADER Regulatory Requirements
PADER requirements for approved drug applications are set out in 21 CFR 314.80(c)(2). The applicant must submit periodic adverse drug experience reports to FDA after approval of the application.
The main requirements include:
- reporting adverse drug experiences that were not already submitted as 15-day Alert reports;
- including a narrative summary and analysis of the information in the report;
- analysing the 15-day Alert reports submitted during the reporting interval;
- including the required individual case safety reports and an index of those cases;
- providing a history of actions taken because of adverse drug experiences, such as labelling changes or studies initiated.
PADERs are submitted quarterly for the first 3 years after approval and annually thereafter, unless FDA requires a different reporting schedule. The detailed due dates are discussed in the submission-timeline section below.
For electronic submissions, FDA’s current eCTD technical guidance states that periodic safety reports have a descriptive portion and individual case safety reports (ICSRs). Only the descriptive portion is submitted in the eCTD, and it may be provided as a PADER in Section 5.3.6 as an individual PDF file. The ICSRs are handled separately and should not be included as E2B XML files within the eCTD sequence.
Components Of A PADER Report
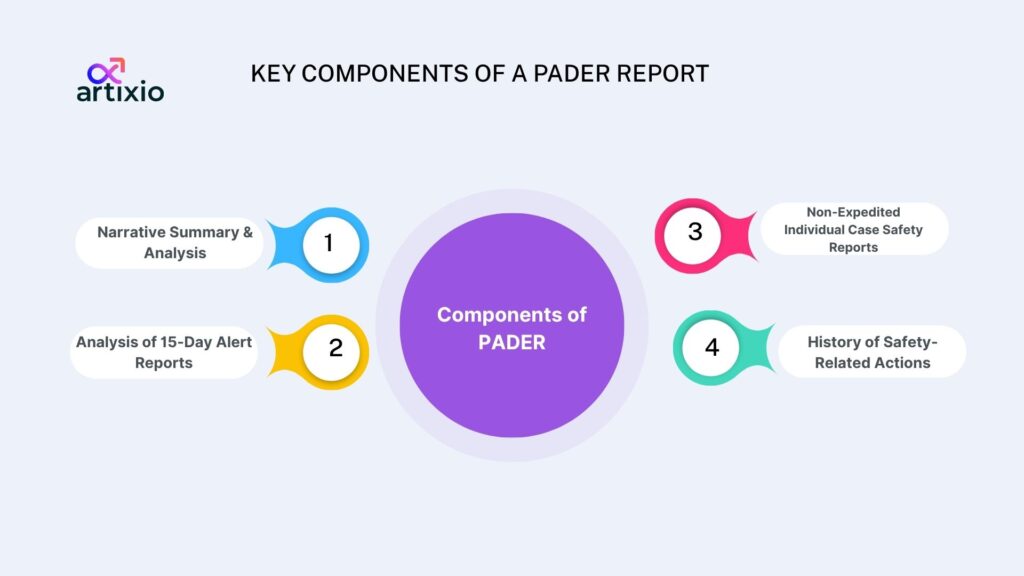
Under 21 CFR 314.80(c)(2), a periodic adverse drug experience report includes the following key information:
1. Narrative Summary and Analysis
The report should provide a narrative summary and analysis of the adverse drug experience information covered during the reporting interval.
2. Analysis of 15-Day Alert Reports
PADER should include an analysis of the 15-day Alert reports already submitted during the reporting period. These reports should be appropriately referenced using information such as the applicant’s patient identification number, adverse reaction term, and date of submission to FDA.
3. Non-Expedited Individual Case Safety Reports
Adverse drug experiences that were not reported as 15-day Alert reports are included as non-expedited ICSRs associated with the periodic report. An index or line listing of the relevant cases is also required.
4. History of Safety-Related Actions
The report should include actions taken since the previous periodic report because of adverse drug experiences. Examples given in the regulation include labelling changes or studies initiated.
For electronic reporting, FDA separates the descriptive portion of the periodic safety report from the ICSRs. The descriptive PADER may be submitted in eCTD Section 5.3.6 as an individual PDF file, while the associated ICSRs are submitted separately through FDA’s electronic adverse-event reporting system.
FDA accepted E2B(R2) for postmarketing ICSRs through 30 September 2026. From 1 October 2026, postmarketing ICSRs submitted through ESG NextGen must use the E2B(R3) standard.
How To Prepare A PADER?
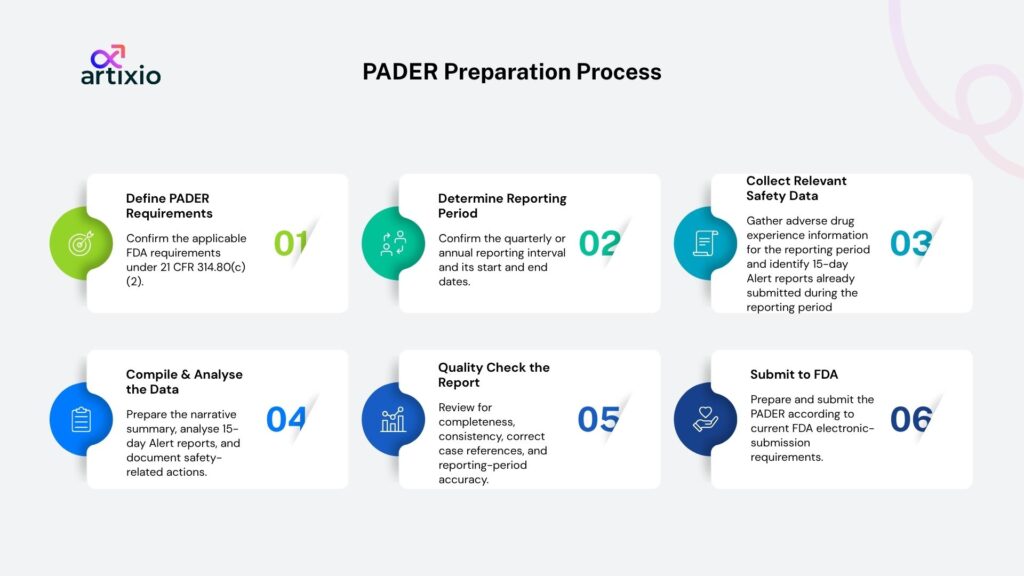
FDA does not prescribe one fixed internal authoring workflow for preparing a PADER. However, the report must contain the information required under 21 CFR 314.80(c)(2). A practical preparation process can include the following steps:
1. Define the reporting period
Confirm whether the PADER is being prepared for a quarterly or annual reporting interval and identify the start and end dates for the period.
2. Collect the relevant adverse drug experience data
Gather the adverse drug experiences that fall within the reporting period. Serious and unexpected cases that meet expedited-reporting requirements should already have been submitted separately as 15-day Alert reports.
3. Review the 15-day Alert reports
Identify the 15-day Alert reports submitted during the interval and prepare the required analysis for inclusion in the periodic report.
4. Prepare the non-expedited case information
Compile the adverse drug experiences that were not submitted as 15-day Alert reports and prepare the required individual case information and case index.
5. Prepare the narrative summary and analysis
Summarize the adverse drug experience information from the reporting period and provide the analysis required by FDA. The summary should be consistent with the underlying case information and the 15-day Alert reports submitted during the interval.
6. Document safety-related actions
Include any actions taken since the previous periodic report because of adverse drug experiences, such as labelling changes or studies initiated.
7. Quality review before submission
Review the report before submission to make sure the case details, references, reporting period, and required information are correct and consistent. Post-approval individual case safety reports should also follow the current ICH E2D(R1) guidance adopted by FDA in March 2026.
8. Prepare the report for electronic submission
The descriptive portion of the periodic safety report should be prepared according to the current FDA electronic-submission requirements and applicable eCTD specifications. FDA continues to update its eCTD technical standards, so applicants should use the currently supported specifications when preparing the submission.
PADER Submission Timelines and Frequency
Under 21 CFR 314.80(c)(2), PADERs are submitted:
- Quarterly for the first 3 years after approval
- Annually after the first 3 years
Each quarterly report is due within 30 days after the close of the quarter. The first quarter begins on the date of approval of the application. Annual PADERs are due within 60 days of the anniversary date of approval.
FDA may require a different reporting schedule in some cases. For example, the agency may extend or re-establish quarterly reporting or require reports at different times through written notice.
The reporting dates should be calculated from the approval date of the application. Current FDA approval letters continue to state that periodic adverse drug experience reporting due dates under 21 CFR 314.80(c)(2) are calculated from the date of approval.
Challenges in PADER Preparation
Preparing a PADER can be difficult when safety data are incomplete, inconsistent, or stored in different places. Applicants also need clear procedures for collecting, reviewing, and reporting postmarketing adverse drug experiences so that the information used in the PADER is accurate and complete.
Common challenges include:
Managing large volumes of safety data
Applicants may need to review substantial amounts of postmarketing safety information during the reporting period. FDA has noted that both industry and regulators face challenges with the timely and efficient collection, processing, and evaluation of individual and aggregate safety data as case volumes increase.
Keeping case information complete and consistent
The PADER summary, 15-day Alert analysis, non-expedited ICSRs, and case index need to align with one another. Missing case information, incorrect references, or inconsistent data can create problems during preparation and review. The regulation requires these elements to be included in the periodic report.
Separating expedited and non-expedited reports correctly
Serious and unexpected adverse drug experiences that meet expedited-reporting criteria must be submitted as 15-day Alert reports. Other reportable adverse drug experiences are handled through the periodic reporting process. Correct classification is therefore important.
Tracking safety-related actions
Applicants must keep a record of actions taken because of adverse drug experiences, such as labelling changes or studies initiated, and include that history in the periodic report.
Meeting reporting timelines
PADERs have defined quarterly and annual due dates. Delays in collecting, reviewing, or reconciling safety information can make it harder to submit the report on time. FDA has taken enforcement action in the past when required periodic reports were not submitted.
Keeping procedures and electronic reporting processes current
FDA periodically updates technical standards and guidance for postmarketing safety reporting. Companies therefore need to keep their internal procedures, case-processing systems, and submission workflows aligned with the current requirements.
PADER vs PSUR vs PBRER
PADER, PSUR, and PBRER are all periodic safety reports, but they are not the same.
PADER is the U.S. FDA report used for approved drug applications under 21 CFR 314.80(c)(2). It includes adverse drug experience information, analysis of 15-day Alert reports, non-expedited case information, and actions taken because of adverse drug experiences.
PSUR, or Periodic Safety Update Report, is the older ICH E2C periodic safety reporting format. It was mainly used to provide periodic safety updates on marketed products.
PBRER, or Periodic Benefit-Risk Evaluation Report, comes from ICH E2C(R2). It developed from the older PSUR format and includes a broader review of cumulative safety information and the product’s benefit-risk balance.
FDA permits a PBRER to be used instead of a PADER when the applicable FDA conditions are met. FDA guidance explains how this alternative format can be used for periodic postmarketing safety reporting.
| Report | Main framework | Main focus | FDA use |
| PADER | U.S. FDA / 21 CFR 314.80(c)(2) | Periodic adverse drug experience reporting | Standard U.S. periodic format |
| PSUR | Older ICH E2C format | Periodic safety update | May be submitted under an approved FDA waiver |
| PBRER | ICH E2C(R2) | Cumulative safety and benefit-risk evaluation | May be submitted instead of PADER under specified FDA conditions |
Conclusion
PADER remains an important part of U.S. postmarketing safety reporting for approved drug applications under 21 CFR 314.80(c)(2). It brings together periodic adverse drug experience information, analysis of 15-day Alert reports, non-expedited case information, and actions taken because of safety findings.
FDA also permits PBRER to be used instead of PADER when the relevant conditions are met. Companies should check which reporting format and schedule apply to their approved application and follow the current FDA requirements for postmarketing safety reporting.
For support with PADER preparation, authoring, review, publishing, submission, and other pharmacovigilance activities, explore Artixio’s Pharmacovigilance Services.
FAQs
Q. What does PADER stand for in pharmacovigilance?
PADER means Periodic Adverse Drug Experience Report. It is a U.S. FDA report used for periodic postmarketing safety reporting under 21 CFR 314.80(c)(2).
Q. Who is responsible for submitting a PADER?
The holder of the approved drug application must submit the required periodic adverse drug experience reports to FDA.
Q. What happens if a PADER does not meet FDA requirements?
If the report is incomplete or does not meet FDA requirements, the applicant may need to correct it or provide more information. Missing deadlines or other reporting requirements can also lead to compliance issues.
Q. Can a PBRER be submitted instead of a PADER?
Yes. FDA allows applicants, under specified conditions, to use the ICH E2C(R2) Periodic Benefit-Risk Evaluation Report (PBRER) in place of a PADER to meet periodic postmarketing safety-reporting requirements.
Q. How often is a PADER submitted?
PADERs are submitted quarterly for the first 3 years after approval and annually thereafter, unless FDA requires a different schedule. Quarterly reports are due within 30 days after the close of the quarter, while annual reports are due within 60 days of the approval anniversary date.
Q. Are PADER and PSUR the same?
No. PADER is the U.S. FDA periodic adverse drug experience report under 21 CFR 314.80(c)(2). PSUR is the older ICH periodic safety update format, while PBRER is the current ICH E2C(R2) periodic benefit-risk format. FDA guidance allows PBRER to be used instead of PADER under specified conditions.